Request a campaign Computational Hit Discovery ServiceA starting point for your target
A ranked, synthetically accessible shortlist of selected molecules delivered in 15–20 daysMolinTimeCase study
What is MolinTime?
An end-to-end in silico hit discovery service that delivers a ranked shortlist of compounds, tailored to your target and screened for predicted binding affinity, ADMET profile and novelty, while remaining within synthetically accessible chemical space.
Hit discovery on your target
Our hit discovery service starts from your target and screens vast variety of the chemical spaces
How we work
Hit Identification
We screen your chosen vast chemical space with a multiparametric pipeline and return a ranked initial shortlist
Structure-based methods to pose and score compounds against the binding site, and evaluate stability & selectivity
Ligand-based methods for affinity & selectivity prediction
Novelty assessment against patents, scientific literature & public databases
In silico ADMET evaluation for solubility, permeability, metabolic stability & toxicity
Hit Expansion
From confirmed hits, we generate new analogues using reaction-based methods and Matched Molecular Pairs (MMP)-based approaches
Every generated molecule is restricted to synthetically accessible chemical space and scored with the same multiparametric pipeline used for hit identification
What you receive
A ranked list of compounds, prioritised for synthesis and biological testing
Case Study
Pikaïros computational hit design paired with Enamine synthesis & experimental validation.
Target implicated in inflammatory, autoimmune, and oncological disease.
Hit Identification
Starting from Enamine’s in-stock collection (~4 M compounds)
~4 M
compounds in the screened collection
160
compounds were computationally preselected and experimentally assayed
1
confirmed hit at 230 nM (0.23 µM)
Hit Expansion
We generated new analogues within Enamine REAL Space and scored them using the same pipeline:
174
compounds tested
71
with >50% inhibition at 20 µM
45
with >70% inhibition at 20 µM
40
compounds selected for dose-response
16
of 40 confirmed as submicromolar
40 compounds selected for dose-response
Best lead
35 nM
next best 81 nM
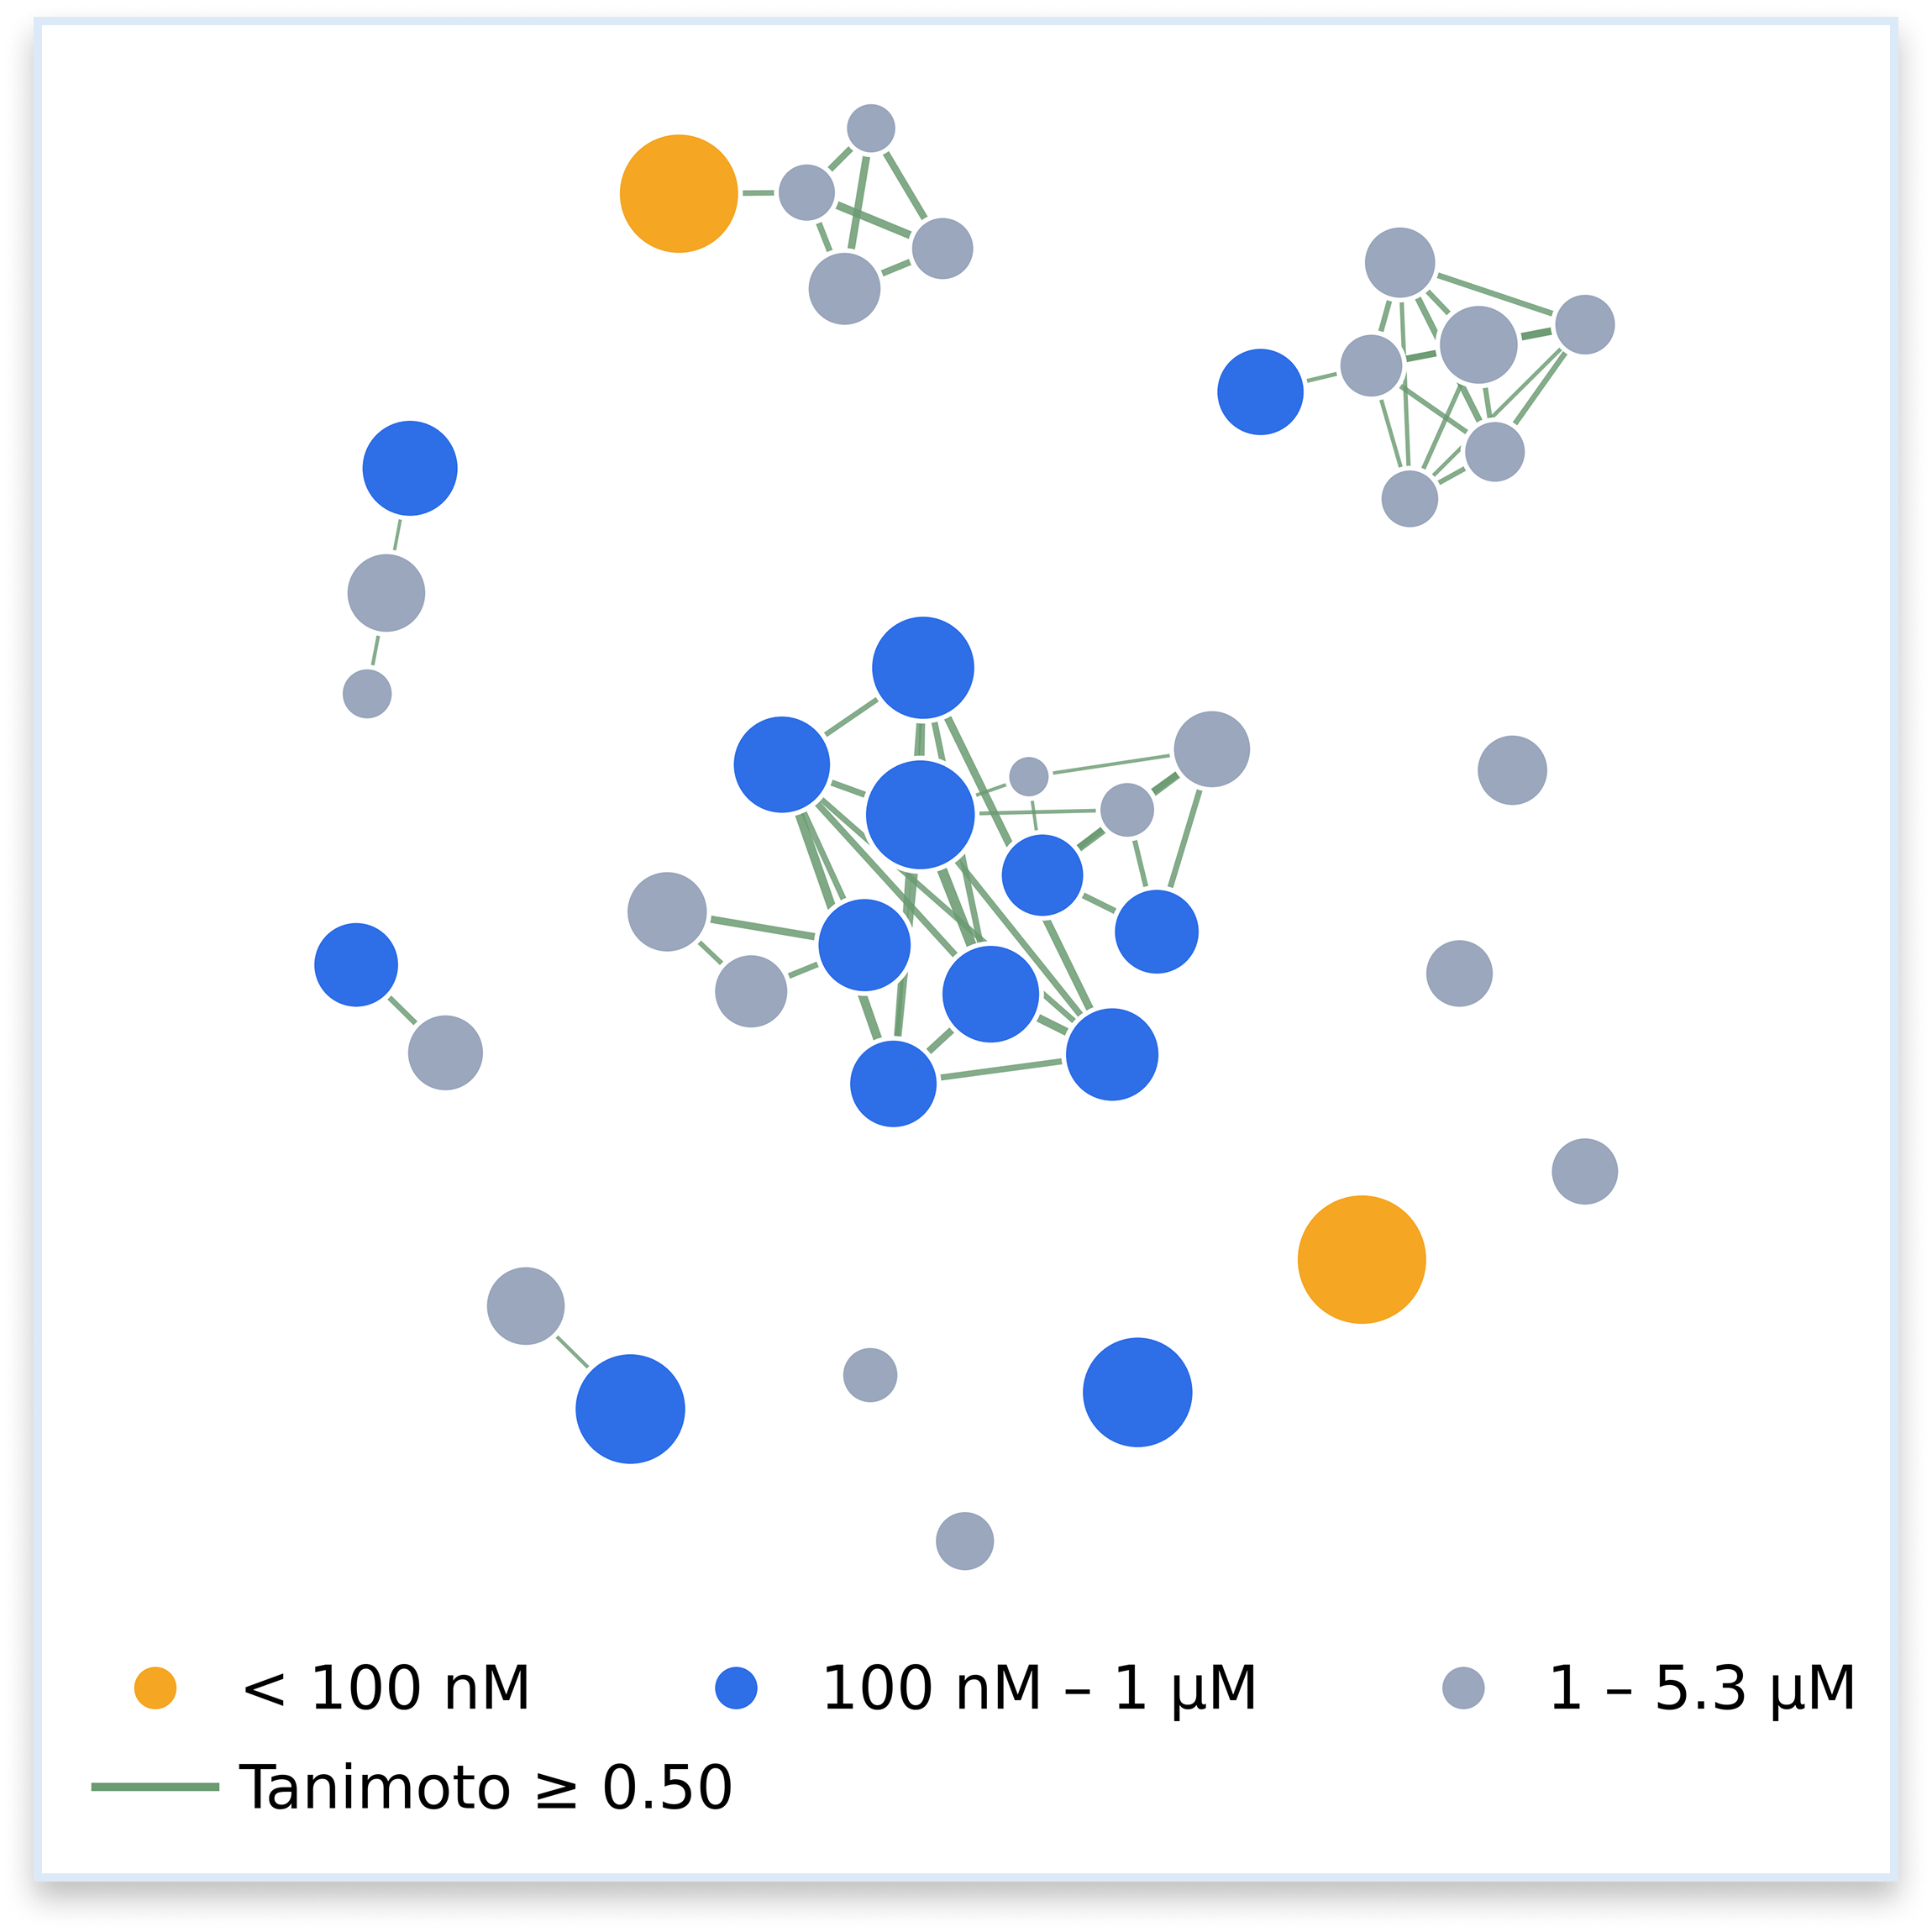
IC₅₀ range 35 nM – 5.3 µM (median ~1.2 µM)
16 of 40 confirmed as submicromolar
Best lead 35 nM, next best 81 nM
Across the 174 selected molecules, mean nearest-neighbour ECFP4 Tanimoto similarity was 0.38 to ChEMBL 37 and 0.35 to target-associated compounds drawn from public databases and patents
Nearest-neighbour similarity* to target-associated compounds (full molecule)
Mean 0.35 · Max 0.46
Nearest-neighbour similarity* to target-associated compounds (scaffold)
Mean 0.38 · Max 0.55
Structural diversity
146 of 174 unique Murcko scaffolds
*ECFP4 Tanimoto, 2048-bit
Tailored Chemical Collections
Each library is built around a single drug target and delivers a ranked, annotated set of molecules, from hundreds to thousands per collection, generated through machine learning and docking. Every molecule is sourced from commercial catalogues, with predicted potency, selectivity, and ADMET profile already assessed, so it is ready for synthesis and experimental screening without further preprocessing.
Target biology, disease context, known chemical space, and available structural data are assessed to define a concrete set of hit-identification objectives.
02
Database & patent curation
Public chemical and biological databases are mined alongside patent landscapes to map existing chemical matter. Novelty is benchmarked against this auditable reference set, ensuring each delivered molecule is potentially novel relative to a defined baseline.
03
Chemical space exploration
Novel candidates are generated through ML-driven ligand-based and structure-based methods to identify molecules with strong potential to be active against the target of interest.
04
Multi-criteria prioritization
Candidates are scored across novelty, predicted potency, ADMET profile, and synthetic accessibility. Compounds flagged for hERG inhibition, structural alerts, PAINS, or target-specific liabilities are excluded. Models are validated against held-out experimental data.
05
Output
A ranked collection of hundreds to thousands of structurally diverse molecules, each sourced from commercial vendor catalogues with supplier name, indicative pricing, and lead time. Here, synthetic accessibility means a compound ready to order and assay.
We use cookies to ensure that we give you the best experience on our website. If you continue to use this site we will assume that you are happy with it.